Category: publications
| Authors: |
Brent Motmans, Digvijay Ghogare, Thijs G.I. van Wijk, Joren Van Herck, Saba Heidarian, Pieter De Meyer, Berend Smit, An Hardy, and Danny E. P. Vanpoucke |
| Journal: |
Chem. Mater XX, YYY (2026) |
| doi: |
ZZ |
| IF(2025): |
7.1 |
| export: |
bibtex |
| pdf: |
<ChemMater_XX> <ArXiv> |
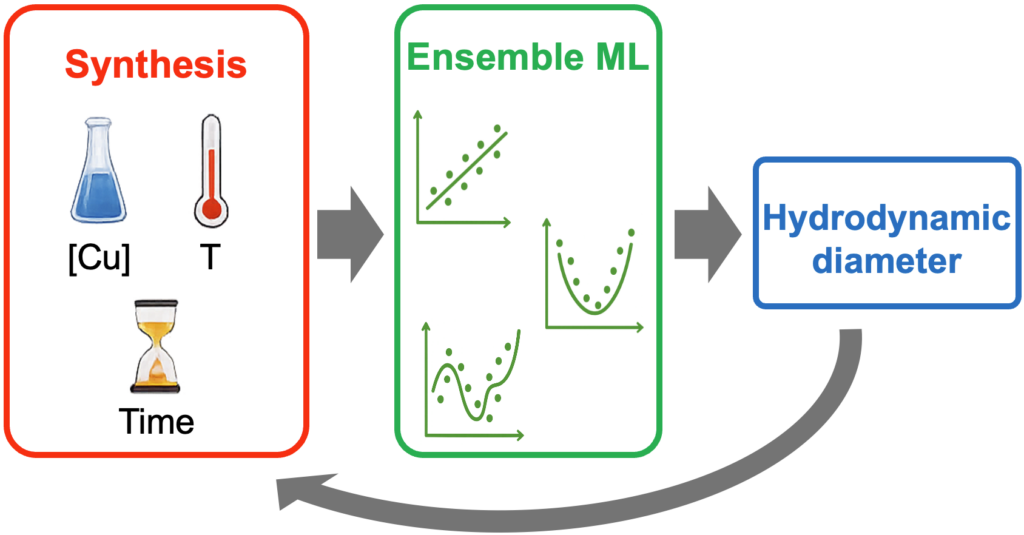 |
| Graphical Abstract: A machine learning approach to predict the hydrodynamic diameter of Cu Nanoparticles using a small data approach. |
Copper nanoparticles (Cu NPs) have a broad applicability, yet their synthesis is sensitive to subtle changes in reaction parameters. This sensitivity, combined with the time- and resource-intensive nature of experimental optimization, poses a major challenge in achieving reproducible and size-controlled synthesis. While Machine Learning (ML) shows promise in materials research, its application is often limited by scarcity of large high-quality experimental data sets. This study explores ML to predict the DLS-derived hydrodynamic diameter of Cu NPs from microwave-assisted polyol synthesis using a small data set of 25 in-house performed syntheses. Latin Hypercube Sampling is used to efficiently cover the parameter space while creating the experimental data set. Ensemble regression models successfully predict hydrodynamic diameters with good predictive performance given the limited dataset. Since quantitative regression requires a unique DLS-derived hydrodynamic diameter, the regression model is restricted to mono-modal DLS distributions, while a complementary classification model identifies synthesis conditions for which quantitative prediction is applicable. Using equivalent out-of-sample validation (OOB and LOOCV), the ML and DoE models showed comparable generalization (MAE -= 40.92 and 40.46 nm, respectively). The final ensemble model achieved an R2 of 0.74, an MAE of 23.81 nm, compared to an R2 of 0.60, an MAE of 33.54 nm for the DoE model, while retaining the complete synthesis parameter space, making it better suited for synthesis guidance. Additionally, classification models using both random forests and Large Language Models (LLMs) are evaluated to distinguish between large and small particles. These classification models exhibited only modest predictive performance, indicating that this small dataset is insufficient to fully exploit the capabilities of complex LLMs. Overall, this study demonstrates that carefully curated small data sets, paired with robust classical ML, can effectively support the synthesis of Cu NPs and highlights that for lab-scale studies, complex models like LLMs may offer limited benefits. The validation experiments further indicate the potential of ML guided synthesis to improve the efficiency of experimental optimization and reduce resource consumption.
Permanent link to this article: https://dannyvanpoucke.be/2026-paper_cunp_mlbrent-en/
| Authors: |
Stijn Lenaers, Stijn Lammar, Anurag Krishna, Brent Motmans, Bart Ruttens, Jan D’Hean, Tom Aernouts, Danny E.P. Vanpoucke, Koen Vandewal, Giuseppe Portale, Laurance Lutsen, Dirk Vanderzande, and Wouter T.M. Van Gompel |
| Journal: |
Sol. Energy Mater. Sol. Cells 307, 114662 (2026) |
| doi: |
10.1016/j.solmat.2026.114662 |
| IF(2025): |
6.6 |
| export: |
bibtex |
| pdf: |
<JSolMat> |
Inverted p-i-n perovskite solar cells (PSCs) have drastically increased in efficiency in recent years, partially due to the inclusion of self-assembling molecules (SAMs) as hole transporting materials (HTLs) and the addition of large organic ammonium salts as a passivating interlayer. In this study, the effect of halogen-functionalized carbazole-based ammonium salts as interlayers between the perovskite absorber and different HTLs is investigated. Fluorinated (F2-Cz), chlorinated (Cl2-Cz), and brominated (Br2-Cz) derivatives are synthesized and incorporated into p-i-n PSCs using NiOx, PTAA, 2PACz, and 4PAPyr as HTLs. All interlayers improve the open-circuit voltage (Voc), indicating effective defect passivation. Notably, a systematic increase in current density (Jsc) and overall PCE is observed across the halogen series from fluorine to bromine. Br2-Cz consistently delivered the highest performance across all tested HTLs, confirming its versatility as an interlayer that is compatible with different types of HTL. The highest power conversion efficiency of 20.9% (0.125 cm2) is achieved when applying the brominated carbazole derivative Br2-Cz on top of the in-house synthesized pyrene-based SAM 4PAPyr. These findings highlight the potential of targeted molecular engineering of interlayers to optimize solar cell performance.
Permanent link to this article: https://dannyvanpoucke.be/2026-paper_carbazole/
| Authors: |
Silviu Florin Acaru, Marc Comí, Panagiotis Falireas, Danny E. P. Vanpoucke, Richard Vendamme, and Katrien Bernaerts |
| Journal: |
Materials & Design 267, 116265 (2026) |
| doi: |
10.1016/j.matdes.2026.116265 |
| IF(2025): |
7.9 |
| export: |
bibtex |
| pdf: |
<Mat&Des_267> |
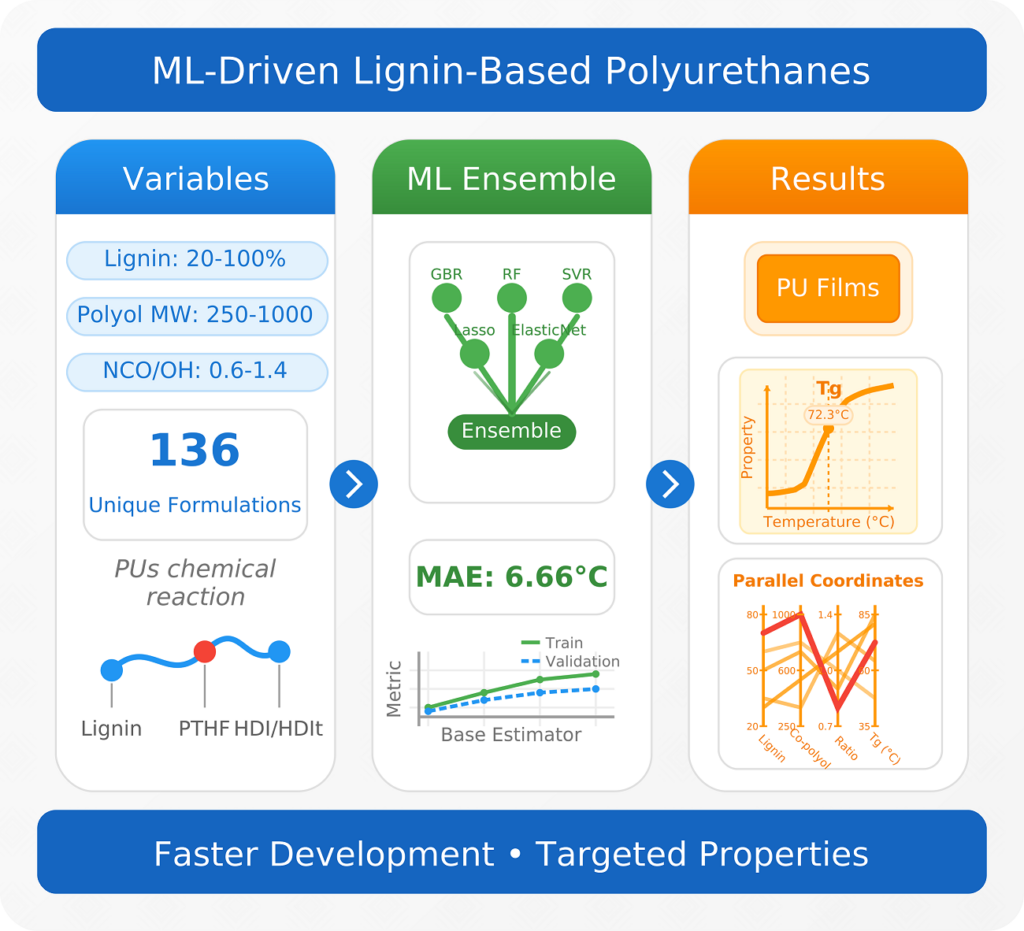 |
| Graphical Abstract: A machine learning ensemble accurately predicts glass transition temperatures for lignin-based polyurethanes within 6.66°C, despite being trained on a dataset of only 136 formulations. The model efficiently combines seven key features to overcome this data limitation. This optimized workflow generated over 4 million novel lignin-polyurethane formulations, with an intuitive interface accelerating the adoption of sustainable polyurethanes. |
Lignin-based polyurethanes (PUs) offer a compelling route towards sustainable material development, yet the challenge of designing chemical formulations with targeted properties, such as glass transition temperature (Tg), remains unresolved. In this work, we present a systematic approach, to explore key structural parameters—such as lignin content, polyol chain length, isocyanate functionality, and mixing ratios—across 136 unique formulations, creating a diverse dataset of ligninbased PUs. By harnessing this small dataset, we develop a machine learning (ML) ensemble model capable of accurately predicting Tg, with a mean absolute error of just 6.66°C on the validation set, surpassing the performance of conventional regression methods. Additionally, we enhance model interpretability by integrating advanced mapping techniques and employ an adaptive grid search algorithm to explore extrapolative scenarios. Our workflow, paired with a user-friendly interface, enables rapid discovery and optimization of formulations with desired properties. This study not only deepens the understanding of structure-property relationships in lignin-PUs but also provides a scalable ML-driven tool for designing sustainable materials with precision, highlighting the transformative potential of artificial intelligence in green chemistry and materials innovation.
Permanent link to this article: https://dannyvanpoucke.be/2026-paper_digiligninsylviu-en/
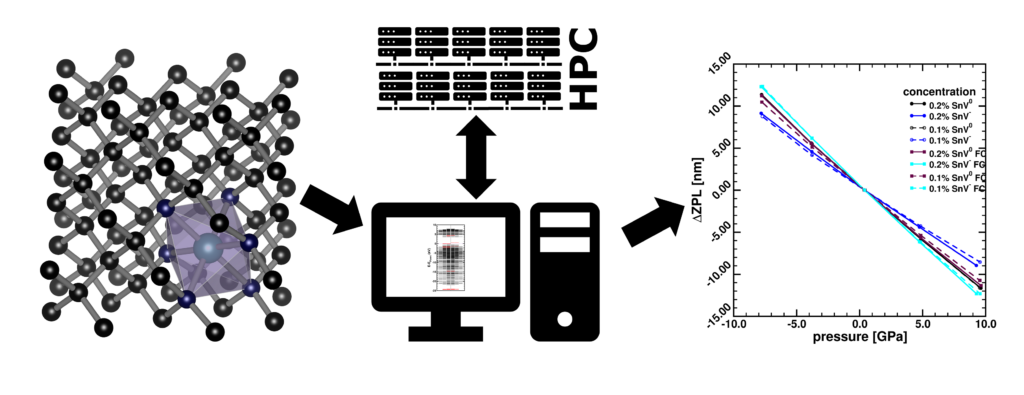 |
| Graphical Abstract: The SnV color center is modelled using first principle calculations to predict the zero-phonon-lines in diamond under hydrostatic strain. |
Among the group-IV vacancy color centers in diamond, the SnV holds promise for photonics based quantum applications. In this work, the Tin-Vacancy (SnV) zero-phonon line (ZPL) and its pressure coefficient are calculated using first principles approaches. The predicted absolute ZPL position is shown to be strongly influenced by the method and supercell size used. The results are therefore extrapolated to the dilute limit allowing for direct comparison with experiments. The importance of identifying the color-center related Kohn–Sham states is highlighted, as well as the shifting of these states due to electron excitations as well as supercell size and k-point position. In contrast to the absolute ZPL positions, the relative position of the SnV0 ZPL is consistently redshifted about 43 nm compared to the SnV– ZPL. In addition, the pressure coefficient is shown to be very robust over different methods, always resulting in a value of about 1.4 nm/GPa, for both SnV0 and SnV–. Finally, the computational accuracy and cost are put into perspective.
Permanent link to this article: https://dannyvanpoucke.be/2026-paper_snvcolorcenter/
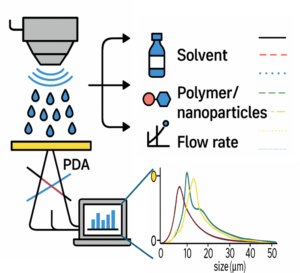 |
| Graphical Abstract: This study explores how machine learning models, trained on small experimental datasets obtained via Phase Doppler Anemometry (PDA), can accurately predict droplet size (D₃₂) in ultrasonic spray coating (USSC). By capturing the influence of ink complexity (solvent, polymer, nanoparticles), power, and flow rate, the model enables precise droplet control paving the way for optimized coatings in advanced functional materials. |
This study examines droplet formation in ultrasonic spray coating (USSC) as a function of ink formulation (solvent, polymer, nanoparticles). First, acetone with polyvinylidene fluoride (PVDF) at concentrations from 0-4.5 wt% is used to examine the effect of polymer additions. Additionally, acetone-based SiO2 nanofluids (0-10 g/L), are explored. Finally, the combination of both polymer (PVDF) and nanoparticles (SiO2) in acetone is studied. Droplet sizes are measured using Phase Doppler Anemometry under varying atomization power and flow rates. Machine Learning (ML) algorithms are employed to develop droplet size models from key spray parameters, including atomization power, flow rate, polymer concentration, and nanoparticle concentration. The model shows significantly higher accuracy than existing empirical models. The model is further validated on IPA-based inks with polyethylenimine (PEIE) or ZnO nanoparticles, and on acetone–cellulose acetate formulations, confirming its robustness across diverse ink systems. In addition to revealing the influence of coating parameters on the droplet formation and distribution, obtained both via experimental validation and ML, this study demonstrates that ML can be effectively applied to small experimental datasets, offering a robust framework for optimizing droplet formation and understanding key spray parameters in USSC for complex, unexplored inks enabling novel coating applications.
Permanent link to this article: https://dannyvanpoucke.be/2025-paper_mldroplets_pieterverding-en/
| Authors: |
Goedele Roos, Danny E.P. Vanpoucke, Ralf Blossey, Marc F. Lensink, and Jane S. Murray |
| Journal: |
J. Chem. Phys. 163, 114112 (2025) |
| doi: |
10.1063/5.0268712 |
| IF(2023): |
3.1 |
| export: |
bibtex |
| pdf: |
<JChemPhys_163> |
 |
| Graphical Abstract: The Electrostatic Potential of water in different situations. On the left two interacting water molecules are shown, while on the right a water molecule interacting with a protein model representation is shown. |
The electrostatic potential plotted on varying contours (VS) of the electron density guides us in the
understanding of how water interactions exactly take place. Water—H2O—is extremely well balanced, having a hydrogen VS,max and an oxygen VS,min of similar magnitude. As such, it has the capacity to donate and accept hydrogen bonds equally well. This has implications for the interactions that water molecules form, which are reviewed here, first in water–small molecule models and then in complex sites as lactose and its crystals and in protein–protein interfaces. Favorable and unfavorable interactions are evaluated from the electrostatic potential plotted on varying contours of the electronic density, allowing these interactions to be readily visualized. As such, with one calculation, all interactions can be analyzed by gradually looking deeper into the electron density envelope and finding the nearly touching contour. Its relation with interaction strength has the electrostatic potential to be used in scoring functions. When properly implemented, we expect this approach to be valuable in modeling and structure validation, avoiding tedious interaction strength calculations. Here, applied to water interactions in a variety of systems, we conclude that all water interactions take the same general form, with water behaving as a “neutral” agent, allowing its interaction partner to determine if it donates or accepts a hydrogen bond, or both, as determined by the highest possible interaction strength(s).
Permanent link to this article: https://dannyvanpoucke.be/2025-paper-wateresp-roos-en/
 |
| Graphical Abstract: Schematic representation of the electrostatic potential within a water molecule along the lines between the atoms. The color background shows the electrostatic potential on the 0.001 a.u. contour of the density. The Vs,min and Vs,max points on the surface are indicated. |
This paper discusses the use of the electrostatic potential in both recent and older literature, with an emphasis upon a 2022 Molecular Physics article by Politzer and Murray entitled “Atoms do exist in molecules: analysis using electrostatic potentials at nuclei“. We discuss electrostatic potentials at nuclei and how they easily lead to atoms in molecules, without physically separating the individual atoms. We further summarize the work by the Politzer group on definitions of atomic radii by means of the electrostatic potential. The earlier studies began in the 1970’s and continued through the 1990’s. Unfortunately, access to these older publications is often limited, cfr. digital libraries often limit the authorized access until a certain publication year, and these papers are often not cited in current publications. Although still being highly interesting and relevant, this older literature is in danger of being lost. Digging into this older literature thus opens up new views. Our feeling is that Peter passed ‘on’ a vision that boundaries do not exist between atoms in molecules, but that some useful and meaningful radii can be obtained using the electrostatic potential between atoms in molecules.
Permanent link to this article: https://dannyvanpoucke.be/2025-paper-esp-politzer-rev-en/
| Authors: |
Thijs G.I. van Wijk, E. Aylin Melan, Rani Mary Joy, Emerick Y. Guillaume, Paulius Pobedinskas, Ken Haenen, and Danny E.P. Vanpoucke |
| Journal: |
Carbon 234, 119928 (2025) |
| doi: |
10.1016/j.carbon.2024.119928 |
| IF(2024): |
10.5 |
| export: |
bibtex |
| pdf: |
<Carbon> |
 |
| Graphical Abstract: Schematic representation of the impact of hydrostatic and linear strain on the Zero Phonon Line of the neutral GeV defect in diamond. |
Color centers in diamond, such as the GeV center, are promising candidates for quantum-based applications. Here, we investigate the impact of strain on the zero-phonon line (ZPL) position of GeV0. Both hydrostatic and linear strain are modeled using density functional theory for GeV0concentrations of 1.61 % down to 0.10 %. We present qualitative and quantitative differences between the two strain types: for hydrostatic tensile and compressive strain, red- and blue-shifted ZPL positions are expected, respectively, with a linear relation between the ZPL shift and the experienced stress. By calculating the ZPL shift for varying GeV0 concentrations, a shift of 0.15 nm/GPa (0.38 meV/GPa) is obtained at experimentally relevant concentrations using a hybrid functional. In contrast, only red-shifted ZPL are found for tensile and compressive linear strain along the ⟨100⟩ direction. The calculated ZPL shift exceeds that of hydrostatic strain by at least one order of magnitude, with a significant difference between tensile and compressive strains: 3.2 and 4.8 nm/GPa (8.1 and 11.7 meV/GPa), respectively. In addition, a peak broadening is expected
due to the lifted degeneracy of the GeV0 eg state, calculated to be about 6 meV/GPa. These calculated results are placed in perspective with experimental observations, showing values of ZPL shifts and splittings of comparable magnitude.
Permanent link to this article: https://dannyvanpoucke.be/2025-paper-strainedgev-en/
| Authors: |
Asif Iqbal Bhatti, Sandeep Kumar, Catharina Jaeken, Michael Sluydts, Danny E.P. Vanpoucke, and Stefaan Cottenier |
| Journal: |
Journal of Materials Chemistry A 13, 526-539 (2025) |
| doi: |
10.1039/D4TA06603K |
| IF(2024): |
10.7 |
| export: |
bibtex |
| pdf: |
<J.Mat.Chem.A> |
 |
| Graphical Abstract: Schematic representation of the LPS material and the variation of results obtained due to slight changes in settings within a High Throughput workflow. |
High-throughput computational screening has become a powerful tool in materials science for identifying promising candidates for specific applications. However, the effectiveness of these methods relies heavily on the accuracy and appropriateness of the underlying models and assumptions. In this study, we use the popular argyrodite solid-state electrolyte family Li6PS5X (X = Cl, Br, I) as a case study to critically examine key steps in high-throughput workflows and highlight potential pitfalls. We demonstrate some of these pitfalls by highlighting the importance of careful structural considerations, including symmetry breaking and site disorder, and examine the difference between 0 K thermodynamic stability and finite-temperature stability based on temperature-dependent Gibbs free energy calculations. Furthermore, we explore the implications of these findings for the ranking of candidate materials in a mini-throughput study in a search space of isovalent analogs to Li6PS5Cl. As a result of these findings, our work underscores the need for balanced trade-offs between computational efficiency and accuracy in high-throughput screenings, and offers guidance for designing more robust workflows that can better bridge the gap between computational predictions and experimental realities.
Permanent link to this article: https://dannyvanpoucke.be/2025-paper-thedevilinthedetails-en/
| Authors: |
Emerick Y. Guillaume, Danny E. P. Vanpoucke, Rozita Rouzbahani, Luna Pratali Maffei, Matteo Pelucchi, Yoann Olivier, Luc Henrard, & Ken Haenen |
| Journal: |
Carbon 222, 118949 (2024) |
| doi: |
10.1016/j.carbon.2024.118949 |
| IF(2022): |
10.9 |
| export: |
bibtex |
| pdf: |
<Carbon> |
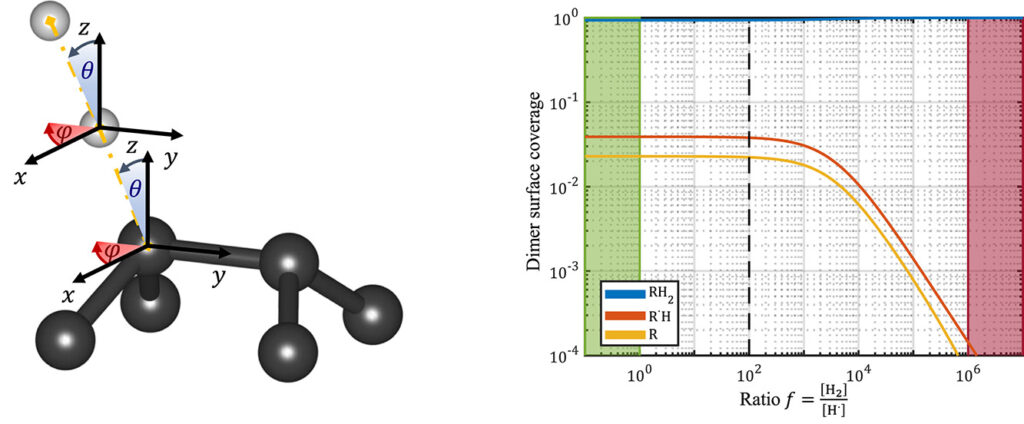 |
| Graphical Abstract: (left) Ball-and-stick representation of aH adsorption/desorption reaction mediated through a H radical. (right) Monte Carlo estimates of the H coverage of the diamond surface at different temperatures based on quantum mechanically determined reaction barriers and reaction rates. |
Hydrogen radical attacks and subsequent hydrogen migrations are considered to play an important role in the atomic-scale mechanisms of diamond chemical vapour deposition growth. We perform a comprehensive analysis of the reactions involving H-radical and vacancies on H-passivated diamond surfaces exposed to hydrogen radical-rich atmosphere. By means of first principles calculations—density functional theory and climbing image nudged elastic band method—transition states related to these mechanisms are identified and characterised. In addition, accurate reaction rates are computed using variational transition state theory. Together, these methods provide—for a broad range of temperatures and hydrogen radical concentrations—a picture of the relative likelihood of the migration or radical attack processes, along with a statistical description of the hydrogen coverage fraction of the (100) H-passivated surface, refining earlier results via a more thorough analysis of the processes at stake. Additionally, the migration of H-vacancy is shown to be anisotropic, and occurring preferentially across the dimer rows of the reconstructed surface. The approach used in this work can be generalised to other crystallographic orientations of diamond surfaces or other semiconductors.
Permanent link to this article: https://dannyvanpoucke.be/2024-paper-hadsorption-emerick-en/